Designing Bulletproof Corrective and Preventive Action (CAPA) Logs for Clinical Operations
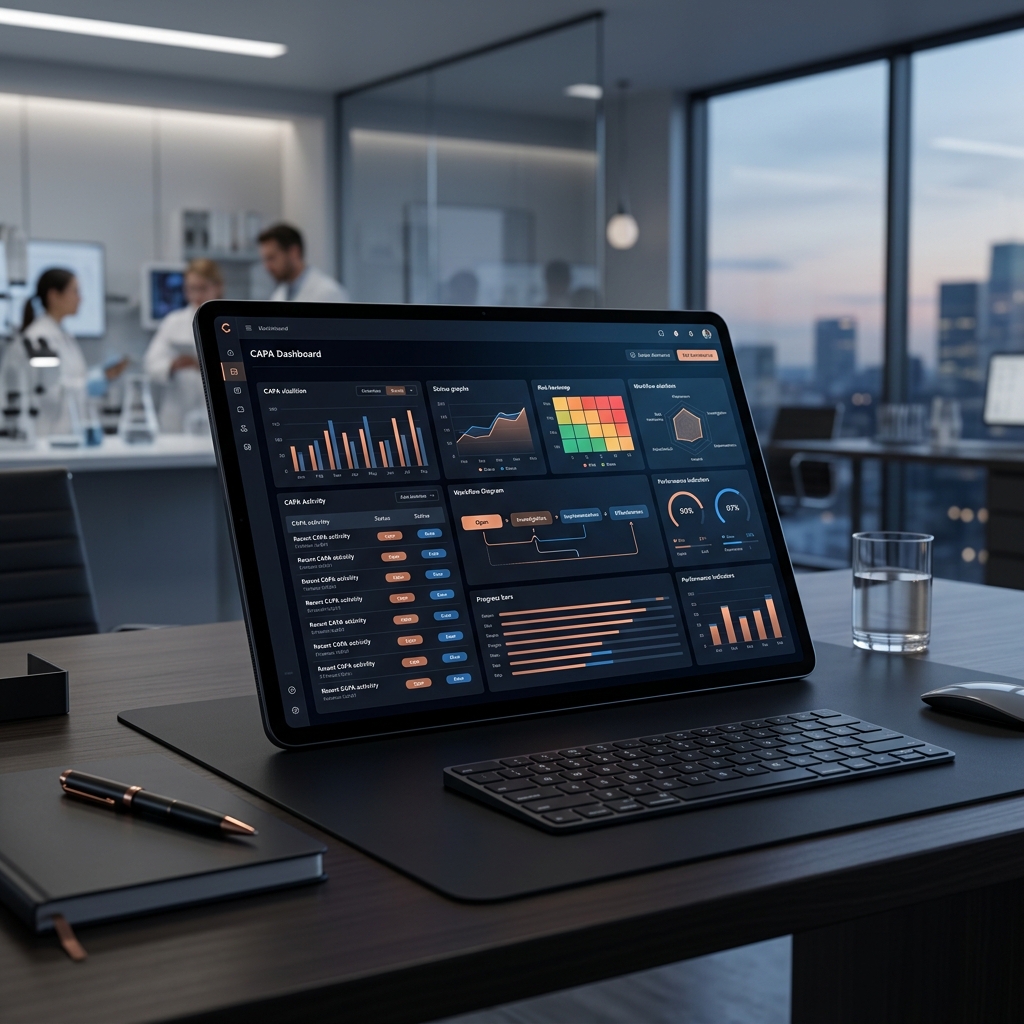
Quick Answer / Executive Summary
- Prioritize Granular Root Cause Analysis: Implement a structured, evidence-based methodology to identify the fundamental source of deviations, moving beyond superficial symptoms to prevent recurrence effectively.
- Enforce ALCOA+ Principles Rigorously: Ensure all CAPA log entries adhere to Attributable, Legible, Contemporaneous, Original, Accurate, and Complete standards to maintain data integrity and withstand regulatory scrutiny.
- Integrate CAPA with Core Operational Logs: Establish explicit linkages between CAPA records and Delegation of Authority (DoA) logs, Quality Tolerance Limits (QTLs), and Trial Master File (TMF) reconciliation efforts to demonstrate holistic quality management.
- Mandate Comprehensive Documentation: Every CAPA entry must detail the non-conformance, immediate action, root cause, corrective action, preventive action, effectiveness check, and closure, supported by verifiable evidence.
- Establish Proactive Effectiveness Checks: Design specific, measurable metrics and timelines for evaluating the sustained impact of preventive actions, validating their efficacy in mitigating future risks.
Designing Bulletproof Corrective and Preventive Action (CAPA) Logs for Clinical Operations
In the highly regulated landscape of clinical research, the meticulous management of quality deviations is not merely a best practice; it is a fundamental requirement for patient safety, data integrity, and regulatory compliance. Corrective and Preventive Action (CAPA) logs serve as the documented backbone of an organization’s Quality Management System (QMS), demonstrating an unwavering commitment to continuous process improvement and the mitigation of future non-conformances. For Clinical Site Directors, Investigators, and CRO Managers, a robust CAPA framework is an indispensable tool for navigating complex operational challenges and preparing for rigorous regulatory inspections. This guide delineates the architectural principles for constructing 'bulletproof' CAPA logs that satisfy regulatory expectations, streamline operational workflows, and fortify the overall integrity of clinical trials.
The Imperative of CAPA in GCP Compliance
Good Clinical Practice (GGCP) mandates a systematic approach to identifying, evaluating, correcting, and preventing deviations from the protocol, standard operating procedures (SOPs), and regulatory requirements. A well-designed CAPA log is the central repository for this critical process, providing an auditable trail of all quality incidents and the measures taken to address them. Regulatory bodies, including the FDA and EMA, routinely scrutinize CAPA systems during site inspections, assessing their effectiveness in safeguarding human subjects and the reliability of study data. A deficient CAPA system can lead to significant findings, consent issues, data anomalies, and, in severe cases, regulatory enforcement actions or disqualification of study data.
Applying ALCOA+ Principles to CAPA Documentation
The integrity of CAPA records is paramount. The ALCOA+ principles, Attributable, Legible, Contemporaneous, Original, Accurate, and Complete, with the added '+' for Consistent, Enduring, and Available, must be the bedrock upon which all CAPA entries are constructed. Each element within the CAPA log, from the initial deviation description to the final effectiveness check, must unequivocally adhere to these standards:
- Attributable: Every action, decision, and observation recorded in the CAPA log must be clearly traceable to the individual who performed it, along with the date and time. This includes identification of the reporter, the individual conducting root cause analysis, and those responsible for implementing actions.
- Legible: All entries, whether electronic or handwritten, must be clear, readable, and unambiguous. Ambiguous language or poorly rendered digital entries undermine the credibility of the record.
- Contemporaneous: Documentation must occur at the time the action is performed or the observation is made. Retrospective documentation can suggest an attempt to fabricate or alter records.
- Original: The first recording of information should be retained, with any subsequent copies clearly identified as such. Raw data supporting the CAPA should be referenced.
- Accurate: Data and information must be truthful and without error. All facts presented must be verifiable against source documentation.
- Complete: The CAPA record must encompass all necessary information required to fully understand the deviation, its impact, the investigation, the actions taken, and the verification of effectiveness. This includes all relevant dates, responsible parties, and references to supporting documents.
Adherence to ALCOA+ transforms a mere log into an irrefutable legal and scientific record.
The Cornerstone: Robust Root Cause Analysis (RCA)
The efficacy of any CAPA hinges on a thorough and accurate Root Cause Analysis (RCA). Superficial identification of symptoms, rather than underlying causes, results in recurring deviations and a perpetual cycle of reactive corrections. CAPA logs must incorporate a structured methodology for RCA, moving beyond the 'what' to uncover the 'why.' Common RCA techniques include:
- 5 Whys: A simple iterative interrogative technique to explore cause-and-effect relationships underlying a particular problem.
- Fishbone Diagram (Ishikawa Diagram): Categorizes potential causes of a problem into main branches (e.g., Man, Machine, Material, Method, Measurement, Environment).
- Failure Mode and Effects Analysis (FMEA): A systematic, proactive method for evaluating a process to identify where and how it might fail and to assess the relative impact of different failures.
The CAPA log must clearly document the RCA methodology employed, the identified root cause(s), and the evidence supporting these conclusions. This section is critical for demonstrating a proactive and analytical approach to quality management.
Operational Workflow Integration: Deviation Reconciliation and Specific Scenarios
Effective CAPA logs do not exist in isolation; they are intricately linked to daily clinical operations and other critical study documentation. The workflow for deviation reconciliation must be seamlessly integrated into the CAPA process.
Delegation of Authority (DoA) Logs
Deviations frequently arise from tasks performed by unqualified or improperly delegated personnel. The CAPA log must include a mechanism to cross-reference non-conformances with the DoA log. For instance, if a study procedure was performed by an individual not explicitly delegated and trained for that task, the CAPA should not only address the immediate deviation but also investigate why the DoA log was circumvented or why training was inadequate, leading to preventive actions related to DoA oversight or training procedures.
Quality Tolerance Limits (QTLs)
ICH E6(R2) introduced the concept of QTLs, defining the acceptable range of variability for critical data or processes. Exceeding a predefined QTL constitutes a significant deviation that necessitates immediate CAPA initiation. The CAPA log for QTL breaches must detail the specific QTL exceeded, the magnitude of the excursion, the impact assessment on subject safety and data integrity, the RCA to understand why the QTL was breached, and the corrective/preventive actions. This demonstrates proactive risk-based quality management.
Temperature Excursions
Management of Investigational Product (IP) and critical laboratory samples requires stringent temperature control. A temperature excursion, even if deemed minor, requires a documented CAPA. The CAPA log entry for a temperature excursion must include:
- Identification of the IP/sample affected.
- Duration and magnitude of the excursion.
- Assessment of impact on product/sample viability (e.g., consultation with sponsor/manufacturer).
- Immediate action taken (e.g., quarantine, relocation).
- RCA (e.g., equipment malfunction, human error, power failure).
- Corrective actions (e.g., equipment repair, recalibration).
- Preventive actions (e.g., enhanced monitoring, backup systems, staff retraining).
- Effectiveness check (e.g., sustained temperature stability post-intervention).
The documentation must reflect a clear understanding of the regulatory requirements for IP handling.
Comprehensive CAPA Log Structure: An Audit Trail Perspective
A 'bulletproof' CAPA log must be structured to provide a clear, chronological, and comprehensive audit trail. Key data fields, consistently applied for every entry, include:
- CAPA ID: Unique identifier for tracking.
- Date Opened: When the deviation was identified.
- Deviation Description: Clear, concise factual statement of the non-conformance.
- Source of Deviation: (e.g., self-identified, monitoring visit, internal audit, regulatory inspection, subject complaint).
- Impact Assessment: Evaluation of potential impact on subject safety, rights, well-being, data integrity, and regulatory compliance.
- Immediate Correction/Containment: Actions taken to mitigate immediate harm or stop recurrence while the full CAPA is developed.
- Root Cause Analysis: Detailed description of the methodology and identified root cause(s), supported by evidence.
- Corrective Action(s): Specific actions taken to eliminate the identified root cause.
- Preventive Action(s): Specific actions taken to prevent recurrence of the deviation or similar deviations.
- Responsible Party & Due Date: For each action item.
- Implementation Date: Actual date each action was completed.
- Effectiveness Check Plan: How and when the effectiveness of the preventive action will be verified (e.g., review of subsequent data, re-audits, process metric monitoring).
- Effectiveness Check Results: Documentation of the outcome of the effectiveness check, confirming sustained mitigation.
- Date Closed & Closed By: When the CAPA was formally closed and by whom, with appropriate sign-off.
- Reference Documents: Links or references to all supporting documentation (e.g., SOPs, training records, monitoring reports, source data).
Each field must be diligently completed, demonstrating an unbroken chain of actions from identification to resolution and verification.
Trial Master File (TMF) Reconciliation and Audit Readiness
The CAPA log is a critical component of the TMF, either as a standalone log or via explicit cross-references within TMF documentation. During TMF reconciliation, all deviations impacting TMF content or processes must be reflected in the CAPA system, and vice versa. An auditor will expect to see a clear link between identified TMF deficiencies (e.g., missing essential documents) and a corresponding CAPA addressing the root cause and preventing future occurrences. The TMF must contain evidence of CAPA implementation and closure, underscoring the interconnectedness of quality documentation. Proactive and thorough TMF reconciliation, particularly focusing on the completeness and accuracy of CAPA records, is a direct pathway to audit readiness.
Conclusion
Designing and maintaining bulletproof CAPA logs is an essential competency for any entity engaged in clinical research. It transcends mere documentation, representing a proactive commitment to quality, patient protection, and scientific integrity. By rigorously applying ALCOA+ principles, conducting meticulous root cause analyses, integrating CAPA into all critical operational workflows, and ensuring comprehensive documentation, Clinical Site Directors, Investigators, and CRO Managers can construct a CAPA system that not only satisfies stringent regulatory requirements but also fosters a culture of continuous improvement. A well-executed CAPA process mitigates risks, enhances operational efficiency, and ultimately, accelerates the delivery of safe and effective medical therapies.
https://www.intelliaiscale.com/compliance-tools
Founder and CEO of IntelliAIScale. Former Clinical Research Coordinator at Hôpital Maisonneuve-Rosemont, CCRP, CCRC, CCRA.