Transitioning from Paper Source Data Gaps to Automated Compliance Workflows
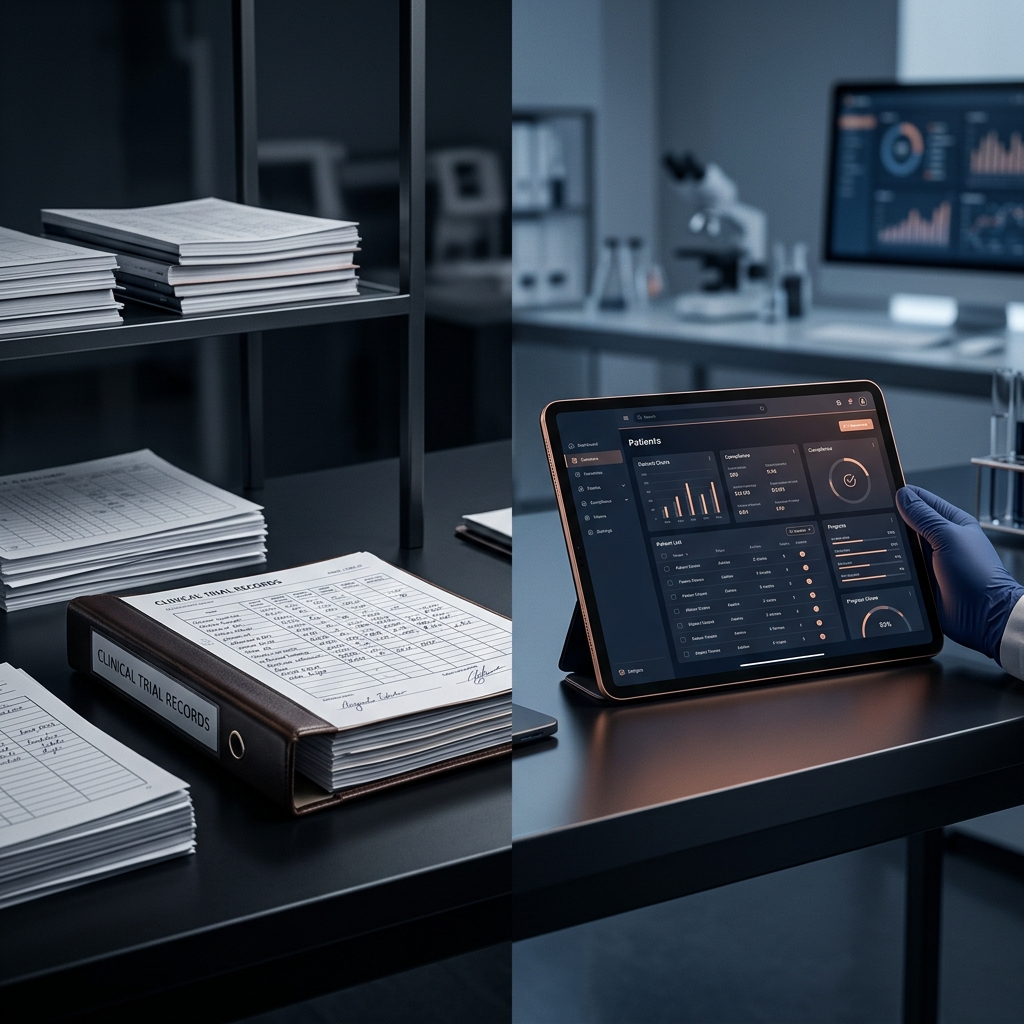
Quick Answer / Executive Summary
- Persistent Risk: Manual paper-based source data perpetuates significant compliance risks including illegibility, transcription errors, incomplete data, and delayed detection of protocol deviations, directly impacting data integrity and regulatory submission reliability.
- Strategic Imperative: Implementing electronic source data (eSource) and automated compliance workflows is no longer optional but a regulatory and operational imperative for maintaining ALCOA+ principles and achieving inspection readiness.
- Key Action Items: Prioritize the digitalization of Delegation of Authority (DoA) logs, integrate real-time Quality Tolerance Limit (QTL) monitoring with automated alerts, deploy IoT solutions for critical environmental data (e.g., temperature excursions), and leverage systems for continuous Trial Master File (TMF) reconciliation.
- Operational Benefit: Automated systems provide verifiable audit trails, enforce protocol adherence, expedite Corrective and Preventive Action (CAPA) initiation and resolution, and significantly reduce the administrative burden associated with manual source data verification.
- Compliance Mandate: Ensure all automated systems adhere strictly to 21 CFR Part 11 requirements for electronic records and signatures, thereby safeguarding the authenticity and integrity of clinical trial data.
Transitioning from Paper Source Data Gaps to Automated Compliance Workflows
Introduction: The Evolving Landscape of Clinical Data Management
The operational landscape of clinical research is undergoing a profound transformation, driven by increasingly stringent regulatory expectations, the imperative for enhanced data quality, and the persistent pressure to accelerate drug development timelines. Central to this evolution is the paradigm shift from traditional paper-based source data documentation to sophisticated automated compliance workflows. For Clinical Site Directors, Investigators, and CRO Managers, understanding and implementing this transition is critical not only for operational efficiency but, more importantly, for maintaining data integrity, ensuring patient safety, and achieving unwavering regulatory compliance. This comprehensive guide delineates the challenges inherent in legacy paper systems and provides a technical roadmap for embracing automated solutions that align with Good Clinical Practice (GCP) and international regulatory standards.
The Pervasive Challenge of Paper Source Data Gaps
Paper source data, while historically foundational, introduces systemic vulnerabilities that directly compromise the reliability and auditability of clinical trial data. Manual transcription from paper records to electronic data capture (EDC) systems is a recognized vector for data entry errors, omissions, and discrepancies. Illegible handwritten entries impede accurate data interpretation and subsequent source data verification (SDV). Furthermore, paper systems inherently lack real-time data accessibility, hindering proactive identification of protocol deviations or safety signals. Version control becomes arduous, leading to the potential for outdated information to persist, and the physical security and archival of paper records present logistical and compliance challenges. These cumulative gaps contribute to an elevated risk profile for a clinical investigation, potentially resulting in regulatory findings during site audits and inspections, necessitating extensive Corrective and Preventive Actions (CAPAs), and ultimately delaying or jeopardizing regulatory submissions.
Foundational Principles: Re-evaluating ALCOA+ in a Digital Context
The ALCOA+ principles, Attributable, Legible, Contemporaneous, Original, Accurate, Complete, Consistent, Enduring, and Available, serve as the bedrock of data integrity in clinical research. While these principles apply universally, their operationalization fundamentally changes with automation. In a paper-based environment, achieving ALCOA+ is often dependent on meticulous manual processes prone to human fallibility. Automated compliance workflows inherently enforce these principles:
- Attributable: Electronic systems with robust access controls and unique user credentials automatically record who performed an action, when, and what the action entailed, eliminating ambiguity.
- Legible: Digital entries inherently guarantee legibility, circumventing issues of handwriting interpretation.
- Contemporaneous: Real-time data entry directly into eSource systems ensures events are recorded as they occur, preventing retrospective documentation discrepancies.
- Original: Data captured directly in a validated eSource system is the original record, obviating the need for transcription.
- Accurate: Built-in validation checks, range checks, and mandatory fields minimize data entry errors and enhance accuracy at the point of capture.
- Complete: Automated systems can enforce mandatory fields and prompt users for missing data, ensuring comprehensive record-keeping.
- Consistent: Standardized data entry forms and terminology enforce consistency across all records.
- Enduring: Digital records, when properly archived and backed up within a validated system, possess superior longevity and resistance to degradation compared to paper.
- Available: Secure, permission-based electronic access ensures data is readily available to authorized personnel, irrespective of geographical location.
Operationalizing Automated Compliance Workflows
Electronic Source Data (eSource) Implementation
The adoption of eSource represents a pivotal shift. Rather than capturing data on paper first and then transcribing it, eSource involves direct data entry into an electronic system at the point of origin. This eliminates transcription errors, provides immediate data validation, and offers real-time visibility into the study's progress and data quality. Successful eSource implementation requires rigorous system validation according to regulatory guidelines, comprehensive user training, and a robust data governance framework. The system must possess immutable audit trails, electronic signature capabilities compliant with 21 CFR Part 11, and secure data storage mechanisms to maintain data integrity and authenticity throughout the trial lifecycle.
Automating FDA Documentation Compliance
Adherence to FDA regulations, particularly 21 CFR Part 11 for electronic records and electronic signatures, is non-negotiable. Automated systems must be designed and validated to meet these requirements. Key functionalities include secure system access, audit trails that capture all data modifications (including original and changed values, timestamps, and user identities), version control for documents and data, and electronic signatures that are equivalent to handwritten signatures. Leveraging validated software solutions ensures that the entire lifecycle of electronic documentation, from creation and approval to archival, is compliant, reducing the burden of manual compliance checks and mitigating regulatory risk.
Streamlining Delegation of Authority (DoA) Logs
Manual Delegation of Authority (DoA) logs are prone to errors, incomplete entries, and outdated information regarding staff qualifications and delegated tasks. Automated DoA systems centralize this critical documentation, allowing for real-time updates and clear visibility into who is authorized to perform specific protocol-required tasks. These systems can integrate with training matrices, verifying staff qualifications before delegation. Automated alerts can notify investigators and site staff of impending credential expirations or mandatory training requirements, ensuring continuous compliance and eliminating the risk of unauthorized personnel performing study procedures. This directly supports the GCP principle that all staff are qualified by education, training, and experience to perform assigned tasks.
Proactive Management of Quality Tolerance Limits (QTLs)
ICH E6(R2) mandates the establishment of Quality Tolerance Limits (QTLs) to identify systematic deviations from quality requirements that might impact patient safety or data integrity. Automated systems facilitate the proactive monitoring of QTLs by integrating data from various sources (e.g., EDC, eTMF, CTMS). Centralized dashboards can display key risk indicators (KRIs) in real-time, providing immediate visual alerts when predefined QTLs are approached or breached. This enables rapid investigation and initiation of CAPAs, transforming QTL management from a retrospective exercise into a continuous, prospective quality assurance process. Such automation minimizes the likelihood of critical quality issues escalating undetected.
Temperature Excursion Protocols and Automated Monitoring
Maintaining the integrity of Investigational Product (IP) and critical study materials often depends on strict temperature control. Manual temperature logging and retrospective review are inefficient and prone to human error, often leading to delayed detection of temperature excursions. Automated monitoring systems, leveraging Internet of Things (IoT) sensors, provide continuous, real-time temperature data logging. These systems can be configured to trigger immediate, multi-channel alerts (e.g., SMS, email) to designated personnel upon deviation from predefined temperature ranges. Integration with inventory management and CAPA systems allows for expedited root cause analysis and appropriate disposition of compromised materials, thereby safeguarding IP integrity and patient safety.
Enhanced Trial Master File (TMF) Reconciliation
The Trial Master File (TMF) serves as the foundational documentation of the clinical trial, providing verifiable evidence of all trial activities. Manual TMF reconciliation is notoriously time-consuming and often results in gaps or inconsistencies. Automated TMF management systems offer intelligent indexing, cross-referencing capabilities, and real-time completeness checks against established TMF specifications. These systems can identify missing or misfiled documents, highlight impending expiration dates for essential documents, and provide a continuous audit trail of all TMF activities. This automation drastically improves TMF completeness, accuracy, and inspection readiness, ensuring that the study documentation consistently reflects the true conduct of the trial.
Leveraging Corrective and Preventive Action (CAPA) Logs for Continuous Improvement
Automated CAPA management systems are indispensable for a robust quality management system. Integrating these systems with eSource, QTL monitoring, and deviation management tools enables the automated initiation of CAPA records upon the identification of a quality issue or protocol deviation. These systems facilitate the systematic documentation of root cause analyses, the development and implementation of corrective actions to address immediate issues, and preventive actions to preclude recurrence. Automated workflows track the status of CAPAs, assign responsibilities, send reminders, and document effectiveness checks. This ensures a closed-loop CAPA process, driving continuous process improvement and enhancing overall trial quality and compliance.
Strategic Advantages and Regulatory Imperatives
Beyond mitigating compliance risks, the transition to automated workflows offers substantial strategic advantages. It improves data quality, reduces the burden of manual source data verification, and accelerates data lock and database closure timelines. The enhanced visibility and control afforded by automated systems lead to more informed decision-making and better resource allocation. From a regulatory perspective, a well-implemented automated compliance framework provides demonstrable evidence of adherence to GCP, strengthens inspection readiness, and instills confidence in the integrity of submitted data. This proactive approach transforms compliance from a reactive burden into a strategic asset.
Conclusion
The clinical research industry stands at an inflection point. The limitations of paper-based source data systems are increasingly unsustainable in an era of complex protocols and heightened regulatory scrutiny. The strategic adoption of automated compliance workflows, encompassing eSource implementation, sophisticated DoA management, real-time QTL monitoring, advanced temperature excursion protocols, and intelligent TMF reconciliation, is paramount. For Clinical Site Directors, Investigators, and CRO Managers, embracing these technologies is not merely about technological advancement; it is about fortifying data integrity, safeguarding patient well-being, and ensuring the unimpeachable credibility of clinical trial results. The path forward demands a commitment to digital transformation, underpinned by a rigorous understanding of regulatory requirements and a focus on operational excellence.
https://www.intelliaiscale.com/compliance-tools
Founder and CEO of IntelliAIScale. Former Clinical Research Coordinator at Hôpital Maisonneuve-Rosemont, CCRP, CCRC, CCRA.